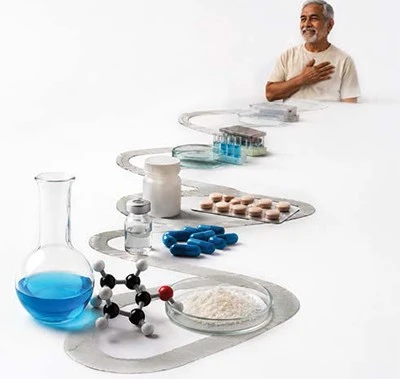
The pharmaceutical industry has never been more effective at developing innovative medicines. Scientific breakthroughs in areas such as gene therapy, oncology and rare diseases are delivering treatment possibilities that would have been inconceivable even a decade ago. Gene therapies are now delivering one-time, potentially curative treatments by introducing functional copies of defective genes, as seen in spinal muscular atrophy and certain inherited retinal diseases. In oncology, highly personalised approaches such as CAR-T cell therapies engineer a patient’s own immune cells to recognise and destroy cancer, requiring complex manufacturing and specialised clinical delivery. At the same time, targeted biologics and precision medicines are being developed for narrowly defined patient populations, often dependent on biomarker testing and coordinated diagnostic infrastructure to ensure appropriate use. For patients living with serious or life-threatening conditions, the announcement of a new therapy can represent a significant moment of hope. However, the journey from scientific breakthrough to real-world treatment is rarely short or straightforward.
The challenge lies in how those innovations are translated into actual patient impact. While the processes that govern regulatory approval and healthcare decision-making remain robust and essential, the complexity of modern therapies means that successful development alone will not enable a seamless transition from regulatory approval into practical, scalable delivery. Innovative therapies today are more targeted, more complex and often more specialised than those that came before. In some cases, existing pathways may not be an option, and a bespoke approach is required to ensure the right clinical infrastructure, supply pathways, healthcare system readiness and local access mechanisms are in place at exactly the right time.
Access Planning as Part of The Development Strategy
Across the industry, there is clear recognition that access is critical. However, the reality is that therapy development has historically been optimised around proving clinical efficacy and safety, while the practical considerations of how a therapy will be delivered, funded and implemented in healthcare systems are often addressed later in the process. As a result, a gap can emerge between regulatory approval and patient access. For example, in Europe, only 28 per cent of all approved treatments are fully avail able and patients now wait an average of 578 days between regulatory approval and access to new medicines (EFPIA, 2026). This delay can have meaningful implications for both organisations and patients. It can limit the realisation of commercial potential and delay reinvestment into future innovation, while also extending the time it takes for patients to access medicines that have already demonstrated clear clinical value.
Bridging this gap is therefore not simply a question of accelerating timelines, but of more closely aligning how therapies are developed with how they are ultimately delivered in practice, requiring a more integrated and strategic approach to access.
One of the key considerations is the robustness of the evidence strategy. Clinical development is rightly designed to demonstrate safety, efficacy and quality to a high regulatory standard. However, the evidence that supports approval does not always satisfy the same evidence criteria required by payers, health technology assessment bodies, clinicians or healthcare providers when deciding how a medicine should be funded, adopted and used. Questions around comparative value, real-world outcomes, long-term durability, budget impact, patient identification and service capacity that are central to access discussions may not be fully addressed if they are considered only after the pivotal development program is already complete.
There is also an operational gap. Due to their complexity and highly personalised nature, many advanced therapies cannot be prescribed and dispensed through established pathways. They may require specialist treatment centres, trained clinical teams, different ordering platforms, diagnostic testing, complex manufacturing slots, cold-chain logistics, individual patient coordination or country-specific import. For some therapies, the practical requirements for delivery can be as important as the clinical profile itself. If those requirements are not recognised and planned for early, companies may find that a product is scientifically ready before the surrounding delivery model is sufficiently enabled.
Geographic Variation Necessitates Careful Planning
Innovation may be developed to address the global unmet need, but access is secured locally. Different countries have different regulatory routes, reimbursement frameworks, clinical practices, supply requirements and levels of healthcare infrastructure. This is particularly relevant in regions such as Asia-Pacific, where markets can be highly diverse in terms of access pathways, funding models and local delivery capabilities. For example, in Korea, a Named Patient Program (NPP) is enabled solely through a dedicated governmental organisation. In Japan, early access is often facilitated through a combination of conditional approval pathways and post-marketing evidence generation requirements, reflecting a strong emphasis on real-world validation. In China, pilot zones such as the Hainan Boao Lecheng International Medical Tourism Pilot Zone allow access to unlicensed medicines under tightly controlled conditions, creating a distinct pathway running in parallel to national regulatory approval. In Australia, access to unlicensed therapies can be supported through mechanisms such as the Special Access Scheme and Authorised Prescriber pathway, each with different evidentiary and oversight requirements. A strategy that works in one country will in most cases not translate directly into another, even when the underlying clinical evidence is the same.
For pharmaceutical and biotech companies, the implication is that access planning cannot sit at the edge of development and commercialisation strategy; it needs to inform strategic decisions much earlier.
Established Commercial Infrastructure does not Guarantee Global Coverage
For established pharmaceutical companies, the access challenge is not a lack of organisational capability. In priority launch markets, large companies usually mobilise extensive regulatory, medical, commercial and supply chain infrastructure. They prepare reimbursement submissions, establish specialist logistics, support treatment centres and build the operational model needed for complex therapies. The challenge is that this infrastructure is generally designed around planned commercial launch. It does not always extend to countries where a product is not registered, not reimbursed, or not expected to be launched in the near term, even where there may be patient populations with significant unmet need. Early access programmes, for example, can address that need for patients pre-commercialisation and help companies understand clinician demand, patient identification, local regulatory requirements and practical delivery considerations. These insights can inform launch planning, evidence generation strategies and supply assumptions before significant commercial investment is made.
Following commercialisation, the challenge remains. Patient needs may continue to exist in countries that sit outside launch markets, but supporting access in those markets often requires capabilities that do not typically exist within a standard commercial operating model. Regulatory navigation, import licensing, patient-specific supply, local compliance requirements, pharmacovigilance oversight and specialist logistics for very small and geographically dispersed patient populations can be resource-intensive to manage internally. As a result, many companies utilise specialist access partners with established regulatory, qual ity and distribution infrastructure to administer managed access programmes on their behalf. This allows manufacturers to support appropriate patient access in non-launch markets without building a full commercial, regulatory and logistics presence in every country where demand may arise.
The Challenge of Access Planning for Biotechs
For emerging biotech companies, the challenge is different. Many are established around a single asset, a small portfolio or a specific scientific platform, with teams heavily concentrated in research, clinical development and regulatory affairs. Commercial infrastructure, market access expertise and global distribution capabilities are often limited or entirely absent. In this environment, access planning can appear daunting, because the scale of the task is disproportionate to the resources available.
Bringing a novel medicine from development into routine clinical practice requires decisions on pricing strategy, reimbursement evidence, geographic prioritisation, patient identification, supply networks and local market requirements. Building these capabilities internally demands significant investment, expertise and time, often at a stage when capital remains focused on progressing clinical programmes and reaching key development milestones.
This creates a strategic decision point for many biotechs. Historically, one route has been to license assets, partially or fully, to larger pharmaceutical companies, in exchange for access to established infrastructure, expertise and market reach. Increasingly, however, companies are exploring alternative partnership models that enable them to retain greater ownership of their assets while accessing specialist external capabilities. Through targeted outsourcing, organisations can draw upon regulatory, access, logistics and commercial expertise without having to build every function internally from the outset. The consequences of these decisions can be significant, influencing the value proposition of the therapy, shaping stakeholder confidence and affecting the speed at which innovation reaches patients.
This is one of the reasons launch performance remains difficult, particularly for first-time launches. Industry analysis has shown that only 20–30 per cent of first-time launchers exceed launch expectations, compared with 40–50 per cent of established companies, while 75 per cent of the least successful launchers reported that market access engagement began too late (McKinsey, 2024). More broadly, nearly two-thirds of therapy launches fail to meet pre-launch expectations in their first year (Guidehouse, 2024). These findings highlight the importance of incorporating access, evidence generation, supply planning and market readiness considerations into development strategy from an early stage, ensuring that critical commercial decisions are informed long before a therapy reaches approval.
Access As a Strategic Design Principle
The most successful access strategies are, therefore, those that are considered as part of the therapy’s broader development and commercialisation journey rather than as a separate activity that begins after approval. For biotechs, this may mean identifying the right combination of partners, capabilities, and markets early enough to support sustainable growth while retaining strategic control of their assets. For larger pharmaceutical companies, it may mean complementing established commercial infrastructure with more flexible models that can reach patients beyond traditional launch markets. Ultimately, when access is viewed not as a downstream operational consideration but as a strategic design principle, it can help companies make more informed development and commercial decisions, support more effective launch execution and, most importantly, ensure that patients have a clearer and more predictable path to treatment wherever they live.
Read the full article — it's free
Register with Pharma Focus Asia to unlock expert insights, research articles and in-depth industry analysis.